

NOTICIAS
EU MDR Update: New Harmonised Standards for Medical Devices
The European Commission has published Implementing Decision (EU) 2026/2479, introducing an important update to the harmonised standards supporting Regulation (EU) 2017/745 (MDR).
Clinical evidence for orphan in vitro diagnostic medical devices
This document provides guidance to manufacturers, notified bodies, expert panels, health institutions, and regulators on the clinical evidence requirements for orphan IVDs and for IVDs that have an orphan indication within the meaning of this guidance. The...
Questions and answers for marketing authorisation holders/applicants referral on nitrosamine impurities in human medicinal products
The European Medicines Agency (EMA) has released Revision 24 of its nitrosamine Q&A, introducing several important updates: https://lnkd.in/gAvk967S

TEAM NB Code of Conduct for Notified Bodies vs 5.3
Team-NB members adopted, at the unanimity of the presents, an updated version od the Code of conduct including a new paragraph requiring notified bodies to have a QMS process to investigate suspected certificate falsification or misuse, preserve evidence, take...
GIRP POSITION PAPER: Revision of the EU Guidelines on Good Distribution Practice for Medicinal Products for Human Use
As the European Union prepares to revise its Good Distribution Practice (GDP) Guidelines, GIRP, the European Healthcare Distribution Association, launches today its front-running position paper setting out key priorities of the sector.GIRP suggests that the new GDP...
New guidance document MDCG 2026-6 on Guidance on clinical evidence for orphan IVDs
This document provides guidance to manufacturers, notified bodies, expert panels, health institutions, and regulators on the clinical evidence requirements for orphan IVDs and for IVDs that have an orphan indication within the meaning of this guidance. The guidance is...
AI in pharmacovigilance – EMA update
Industry Stakeholder Platform Operation of EU Pharmacovigilance 20 October 2026 Luis Pinheiro (Data Analytics and Methods Task Force) Julie Durand (Pharmacovigilance Office)
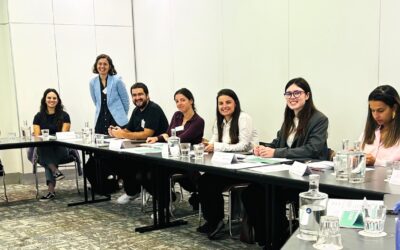
Formação ESTUDOS DE ESTABILIDADE . New ICH Q1 Guideline changes: ” Formação bastante abragente e de alto valor técnico”
A nova ICH Q1 irá impactar em todas as etapas do ciclo de vida do produto; com alterações significativas no desenho das estratégias de estabilidade , condições de armazenamento, interpretação de dados, avaliação estatística dos resuldados, inconsistências / gaps, e...
New revision of the MDCG Position Paper on the management of SS(C)P in Eudamed after mandatory use
1. Current situation in EUDAMED: The summary of safety and clinical performance and the summary of safety and performance (SSCP and SSP respectively), are to be uploaded by the notified body during the process of registering certificates information according to...
PIC/S RECOMMENDATION INTERPRETATION OF THE MOST IMPORTANT CHANGES IN THE ANNEX 1 ON THE MANUFACTURE OF STERILE PRODUCTS
In order to assure a harmonised conduct of inspections, with respect to the 2022 revision of GMP Annex 1, this document provides answers to key questions related to the changes in Annex 1 raised during the revision process and after implementation of the new version....
EMA UPDATES :Notice to sponsors on validation and qualification of computerised systems used in clinical trials
This document, first published on 7 April 2020, has been updated to refer to currently applicable Regulation (EU) No 536/2014 on clinical trials on medicinal products for human use and to the revised ICH E6 (R3) Guideline for Good Clinical Practice. References to...
ISO/TC 215 Task Force 5 WHITE PAPER: Healthcare-Specific Considerations for AI.
Examining how healthcare contexts shape the risks,responsibilities, and evaluation of AI systems and related standards Prepared by ISO/TC 215 Task Force 5 The International Organization for Standardization (ISO) has published a new white paper examining how the...
Six months since implementation of the UK’s new clinical trials regulations
How to avoid delays when submitting clinical trial application assessments The UK's new clinical trial regulations came into force on 28 April 2026, introducing a more flexible and risk-proportionate approach to clinical trial assessment. Sponsors now submit a single...
New policy brief from PHASE IV AI: “EHDS Opportunities for Supporting Research and Innovation”
As the European Health Data Space (EHDS) moves into implementation, how it is put into practice will decide how safely and coherently health data can be used across Europe. Phase4AI-EUProject is a Horizon Europe-funded project advancing the secure and...
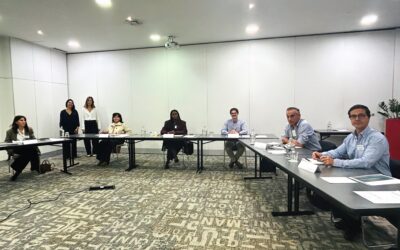
TECH TRANSFER: “Formação completa e robusta “
No passado dia 26 de outubro, decorreu em Lisboa uma nova edição da formação TECH TRANSFER , bajo o lema " A 360° View Across Project Management, Quality, and Regulatory Affairs for Project Success " Esta formação, conduzida por Alina Licova e , Silvia...
FDA Issues Final Guidance on Mandatory Cosmetic Recalls
The U.S. Food and Drug Administration (FDA) issued the final guidance "Questions and Answers Regarding Mandatory Cosmetics Recalls: Guidance for Industry" as part of our ongoing activities to implement the provisions added or required by the Modernization of Cosmetics...
FDA Issues Final Guidance on Prescription Drug Apps and Software
The U.S. Food and Drug Administration today released final guidance establishing a clear framework for how drug companies can develop, distribute, and label apps used alongside their prescription medicines. The guidance ends longstanding uncertainty by providing...

AI Act: Normas harmonizadas para os sistemas de IA de risco elevado
📘 OFFICIAL DOCUMENT
FDA Issues Final Guidance on Prescription Drug Apps and Software
The U.S. Food and Drug Administration today released final guidance establishing a clear framework for how drug companies can develop, distribute, and label apps used alongside their prescription medicines. The guidance ends longstanding uncertainty by...
Plano de Controlo de Qualidade e Conformidade Legal de SUPLEMENTOS ALIMENTARES: ” Formação muito útil para o desempenho das minhas funções “
No passado dia 21 de outubro, decorreu em Lisboa a formação Plano de Controlo de Qualidade e Conformidade Legal de SUPLEMENTOS ALIMENTARES, focada no Controlo de Qualidade, e na Conformidade Legal antes da colocação no Mercado. A formação , conduzida por...
MHRA Registration of medical devices
Guidance on the registration requirements for medical devices placed on the market and put into service Published 20 July 2026
The Use of Data in Public Health. Understanding the Potential for Artificial Intelligence in Medicine
Artificial intelligence (AI) has the ability to transform healthcare for the benefit of both healthcare providers and patients. Implementing a framework that facilitates the (re-)use of data is pivotal to its advancement. However, the development and deployment of AI...
UK Government confirms indefinite recognition of CE marked medical devices
Patients across Great Britain will have continued access to medical devices and in vitro diagnostic devices (IVDs) already approved for the EU market. Patients across Great Britain will continue to have access to the medical devices and in vitro diagnostic...
Methodological Principles for the Assessment of Section 5.1 of the Summary of Product Characteristics
The content of section 5.1 (“Pharmacodynamic properties”) of the Summary of Product Characteristics (SmPC) is proposed by the applicant or marketing authorisation holder and it is assessed by the rapporteur/s. The final content is discussed by the Committee for Human...
Good manufacturing practice (GMP) Multistakeholder workshop on expert contributions to artificial intelligence guidance development (Annex 22)
The objective of this workshop was therefore to gather structured, experience-based input from industry on the control, governance, and mitigation measures applied throughout the AI system lifecycle, with particular emphasis on the application of ICH Q9 quality risk...
PIC/S PE 009-18 – Guide to GMP for Medicinal Products | Part I
Guideline published on: 24/09/2026 Good Manufacturing Practice (GMP) provides the foundation for ensuring that medicinal products are consistently manufactured and controlled according to appropriate quality standards.

EMA Updates Information Page on Bioequivalence Guidelines
On 25 January 2025, the date of coming into effect, ICH Guideline M13A on bioequivalence for immediate-release solid oral dosage forms supersedes applicable parts of this EMA guideline related to bioequivalence study considerations and data analysis for a...
FDA Issues Guidance on Identity and Safety Information for New Dietary Ingredient Notifications
the FDA issued final guidance to help dietary supplement manufacturers and distributors prepare new dietary ingredient notifications (NDINs) with appropriate identity and safety information. The guidance addresses frequently asked questions about NDIN submissions,...
Data Quality Framework for EU Medicines Regulation: Application to the Collection, Management and Submission of Adverse Drug Reaction Reports
This document presents the Adverse Drug Reaction (ADR) chapter of the Data Quality Framework (DQF) for medicines regulation published by the European Medicines Agency (hereafter referred to as European Medicines Regulatory Network (EMRN) DQF). It is an extension...
Clinical Trials Regulation – Q&A Version 7.4
The European Commission has published Version 7.4 of the Questions & Answers on Clinical Trials Regulation (EU) No 536/2014 (CTR), dated October 2026. This document sets out frequently-asked 'questions and answers' regarding the implementation of the rules on...
Clinical trial applications advice pilot will become a permanent initiative from January 2027
The HMA initiative “clinical trial applications advice pilot”, coordinated by the CTCG, will become a permanent initiative from January 2027. This pilot, developed in close collaboration with the European Medicines Agency (EMA), provides clinical trial sponsors with...
Transparency Makes Modern Regulatory Science Stronger — FDA’s Transparency Roadmap
The Transparency Roadmap reflects the agency’s intention to maximize disclosure, guided by what is appropriate and good for public health. It was shaped by soliciting ideas and expertise from innovators across the agency, and it may continue to evolve as we seek the...
The first harmonised standard under the EU AI Act is here
On 14 October 2026, the European Commission published the reference to EN 18286:2026 in the Official Journal of the EU 👉 https://lnkd.in/dkte6BZB With Commission Implementing Decision (EU) 2026/2274, the reference to EN 18286:2026 "Artificial...
PHARMA PACKAGE : Council adopts new rules for a fairer and more competitive EU pharmaceutical sector
The Council has adopted the ‘pharma package’, a new set of rules that will increase patients’ access to medicine and make the EU’s pharmaceutical sector fairer and more competitive. The package represents a far-reaching reform of the EU’s pharmaceutical legislation...
HTA Regulation – Commission selects first medical devices for joint clinical assessment
The Commission has adopted its first decision to select two medical devices for joint clinical assessment under the EU Health Technology Assessment (HTA) Regulation. The two devices are both addressing cardiovascular disease. One of the selected devices is the SAPIEN...
PHARMACOVIGILANCE UPDATE | FDA E2B(R3) IMPLEMENTATION GUIDE — OCTOBER 2026
The FDA has published an updated version of the ICH E2B(R3) Electronic Transmission of Individual Case Safety Reports (ICSRs) Implementation Guide. FDA has adopted the E2B(R3) data standards, and the documents listed below reflect FDA’s recommendations for...
updated PIC/S PI 006-4 guideline RECOMMENDATIONS ON QUALIFICATION AND VALIDATION
The updated PIC/S PI 006-4 guideline replaces the 2007 version (PI 006-3) to harmonize inspection practices with PIC/S GMP Annex 15 and modern lifecycle concepts. It details regulatory expectations across the entire qualification and validation spectrum
Annual report of the Pharmacovigilance Inspectors Working Group for 2025
This document is the sixteenth annual report of the Pharmacovigilance Inspectors Working Group (PhV IWG). The PhV IWG was established by the European Medicines Agency (hereinafter “the Agency”) within the scope of Article 57(1)(i) of Regulation (EC) No 726/2004....
European Medicines Agency post-authorisation procedural advice for users of the centralised procedure
These questions and answers have been produced for guidance only and should be read in conjunction with the rules governing medicinal products in the European Union, volume 2, notice to applicants. MAHs must in all cases comply with the requirements of Community...
ACT EU workplan 2026 – 2027
Technical and regulatory support for preparing clinical trial applications will continue from January 2027 Objectives: • Engage stakeholders, patients • Strengthen EU competitiveness • Enable innovation • Facilitate inclusive clinical trials • Foster excellent...

Advice on ‘end-of-year’ submission dates for type I variations in 2026
Marketing authorisation holders wishing to receive an outcome in 2026 should submit type IA annual updates, individual type IA and type IAIN variations, and super-groupings no later than 20 November 2026. This deadline allows EMA to acknowledge the validity of...
EMA UPDATE : COMPLIANCE with EU GMP Annex 22 and Artificial Intelligence Standards
On october 2 , the EMA published its multi-stakeholder workshop report on Annex 22 ("Artificial Intelligence" ) EMA aims to achieve the following objectives via this workshop: Collect expert insights on responsible AI requirements, risk identification and...
FDA GUIDANCE DOCUMENT – E2B(R3) Electronic Transmission of Individual Case Safety Reports Implementation Guide — Data Elements and Message Specification;
FDA has adopted the E2B(R3) data standards, and the documents listed below reflect FDA’s recommendations for implementing the E2B(R3) data standards. The main document is the E2B(R3) Electronic Transmission of Individual Case Safety Reports (ICSRs) Implementation...
The Council of the European Union has formally adopted the Pharmaceutical Package
The Council has adopted the ‘pharma package’, a new set of rules that will increase patients’ access to medicine and make the EU’s pharmaceutical sector fairer and more competitive. The package represents a far-reaching reform of the EU’s pharmaceutical...
Sponsor Guidance: New Safety Module on the Annual Safety Report
The present document provides CT sponsors with the operational guidance they need to submit ASRs through the New Safety Module on ASR, including responding to the relevant Requests for Information (RFI). It is foreseen that this module will be integrated in the...
A Formiventos está novamente no ranking das Top 5% melhores PME de Portugal
É com grande satisfação que anunciamos que a Formiventos é, pelo quinto ano consecutivo, empresa certificada TOP 5% MELHORES PME PORTUGAL, edição de 2026, certificação atribuída pela SCORING, em termos de desempenho e solidez financeira. Este feito é motivo de...
Electronic Submission Template for Premarket Approval Applications (PMAs)
This guidance describes the technical standards associated with preparation of the electronic submission template for PMAs/PMA supplements that enable submission of the Designated Application Types solely in electronic format. This does not include PMA Modules and...
MDSAP Technical Review and Recognition Decision Procedure
The Medical Device Single Audit Program (MDSAP) has released an updated version of its Technical Review and Recognition Decision Procedure (MDSAP AS P0017.005), effective 1 September 2026. This procedure applies to a technical review process that is to occur...
Essential Principles and Content of Predetermined Change Control Plans. Final Document
The purpose of this document is to share high-level principles on the use of PCCPs as a way of authorizing certain planned medical device software modifications for which regulatory authorization is otherwise required. Additionally, it aims to identify the...
Report on the International Active Pharmaceutical Ingredient Inspection Programme 2017 – 2024
This report covers the period from 2017 to 2024, during which the programme expanded from 10 to 13 authorities, including Health Canada, PMDA Japan, and ANVISA Brazil. The COVID-19 pandemic significantly disrupted on-site inspections activities but also accelerated...
GMP and GDP Question and Answer Guide Version 03 published by ECA
The GMP & GDP Question and Answer Guide has now been published by the ECA in a fully revised Version 03. The guide summarises the Q&As developed by the FDA, EMA/EU, ICH and MHRA, and supplements these with the ECA’s own GMP and GDP Q&As.