

NOTICIAS
Guideline on good pharmacovigilance practices (GVP) Module III – Pharmacovigilance inspections (Rev 2)
This Module contains guidance on the planning, conduct, reporting and follow-up of pharmacovigilance inspections in the EU and outlines the role of the different parties involved. General guidance is provided under III.B., while III.C. covers the overall operation of...
Publication of COMBINE Project 2 Sponsors’ guide on safety reporting in combined studies
This guidance was designed to guide the sponsor on safety reporting from study planning and forecasting of anticipated adverse events to the detection of events during the study conduct, the analysis of causality and seriousness, as well as communication of this...
UK National Commission into the Regulation of AI in Healthcare
The National Commission into the Regulation of AI in Healthcare, United Kingdom has delivered a decisive roadmap that recalibrates how artificial intelligence must be governed, through its latest Recommendations for a future regulatory framework

EMA Article 57 Q&A Version 1.15 (9 September 2026)
📆 On 9 September 2026, the European Medicines Agency (EMA) published Version 1.15 of the document Electronic submission of Article 57(2) data Questions & Answers (Q&As) Version 1.15

EMA guidance on Electronic product information (ePI) submission during centralised procedures
On 1 September 2026, the European Medicines Agency issued practical guidance on the submission of electronic product information, or ePI, for centrally authorised medicinal products. The agency noted that submitting ePI is voluntary until the general pharmaceutical...

EU Medical Device Regulation (2017/745) and In Vitro Diagnostic Medical Device Regulation (2017/746) – Impact on international registrations. Middle East and Africa focus
Considerations for a harmonised approach for handling changes arising from the transition to EU MDR and IVDR Looking at the impact on the Middle East and Africa (MEA) region, where CE marking serves as a major regulatory reference supporting domestic registrations,...

IMDRF Document Implementation Report. 1 September 2026
This document provides a report on the status of implementation of all IMDRF technical documents as self-identified by IMDRF members as of the date of publication. In addition to overseeing IMDRF documents, the IMDRF Management Committee also oversees documents...
EU Regulatory News : IVD Classification Rules – MDCG 2020-16 Rev.5
The European Commission’s Medical Device Coordination Group has published MDCG 2020-16 Rev.5 – Guidance on Classification Rules for in vitro Diagnostic Medical Devices under Regulation (EU) 2017/746 (IVDR), dated September 2026. This guidance, relating to the...
Revised version of EU GMP Annex 19: Reference and Retention Samples
In June 2026, the European Commission published a revised version of EU GMP Annex 19 – Reference and Retention Samples, applicable from 24 September 2026. Legal basis for publishing the detailed guidelines: Article 47 of Directive 2001/83/EC on theCommunity...
Evaluation of Therapeutic Equivalence. Guidance for Industry
📆 August 2026 This guidance provides information regarding FDA’s therapeutic equivalence evaluations, including the assignment of therapeutic equivalence codes (or TE codes). As defined in 21 CFR 314.3(b), therapeutic equivalents are approved drug products...
FDA Considerations for the Regulation of Generative Al-Enabled Medical Devices: Discussion Paper and Request for Feedback
This paper is not intended to propose or implement policy changes regarding how CDRH intends to regulate generative AI-enabled devices. This paper is not intended to communicate CDRH’s proposed (or final) regulatory expectations, including its expectations for...
EMA Guidance for industry on implementing Shortage Prevention Plans (SPP)
The European Medicines Agency has issued an updated guideline to assist marketing authorisation holders (MAHs) to create and maintain drug shortage prevention plans (SPPs) for their products. The main objective of the SPPs is to ensure the MAH has appropriate systems...
PIC/S PI 006-4 “Recommendations on Qualification and Validation”
📆 Effective 1 October 2026: This document updates and renames the previous document, PI 006-3 dated 25 September 2007 titled ‘Recommendations on validation master plan, installation and operational qualification, non-sterile process validation, cleaning...

WHO . Draft Guidelines on GCP
The WHO has recently published a draft guideline, including two annexes, on Good Clinical Practice (GCP) for clinical trials involving medicinal products. Comments on these documents may be submitted until 6 September 2026 Access the documens here: Guidelines for Good...

Product Management Services (PMS) – Implementation of International Organization for Standardization (ISO) standards for the identification of medicinal products (IDMP) in Europe
After the release of version 2 of this guideline, important discussions took place which led to significant changes in the information provided. As a result, version 3 of this document has been completely revised, and previous versions should no longer be taken into...

AI Omnibus enters into force
On 27 July 2026, the AI Omnibus enters into force across the EU, bringing extended timelines to administrative simplification. The AI Omnibus, proposed as part of the digital omnibus package on 19 November 2025, delivers a targeted simplification of the AI rulebook...

EU CTR Q&A Version 7.3 is Out
The European Commission and EU agencies have officially released Version 7.3 of the EU Clinical Trials Regulation (CTR) Q&A guidance. You can download the full, updated guidance directly from here

New WHO guide on Artificial intelligence-related health research
This report considers how best to address this growing gap in ethical oversight. The document examines how standards for research ethics may need to evolve to address the risks and benefits of AI-related health research and to integrate emerging ethical principles...

GDP for APIs: APIC Publishes Updated “How to Do” Document
This document is based on the EU Commission Guideline on the principles of Good Distribution Practice of active substances for medicinal products for human and veterinary use and therefore follows the same structure. This APIC document provides guidance on practical...

Team-NB Position Paper :TRANSFER AGREEMENT FOR SURVEILLANCE OF LEGACY DEVICES
TRANSFER AGREEMENT FOR SURVEILLANCE OF LEGACY DEVICES specifying the terms of the transfer of the appropriate surveillance activities according to Article 120 (3e) of Regulation (EU) 2017/7451 in respect of legacy devices covered by a certificate issued in accordance...

MDR Certification Process (including Pre-application, Application and Post Application phases) – Consensus document
The purpose of this consensus document is to describe in detail the pre-application, application processes through which manufacturers may apply to Notified Bodies (NBs) for the certification of medical devices under the regulation (EU) 2017/745 (MDR). The document...

New ISO Guidance for AI-Powered Medical Devices: ISO/TS 24971-2:2026 Published!
This document provides guidance on risks specific to artificial intelligence (AI) and machine learning (ML) and how to apply the risk management process of ISO 14971 to ML-enabled medical devices (MLMD). This document is intended to be used in conjunction with...
Commission publishes guidelines on transparency obligations for providers and deployers of certain AI systems
The European Commission published guidelines to assist providers and deployers of artificial intelligence (AI) systems in meeting the AI Act's transparency obligations, which start to apply on 2 August 2026. Transparency obligations will help people recognise when...
MDCG Position Paper: UDI assignment between manufacturers and distributors
This MDCG Position Paper aims at explaining the point to clarify the situation and avoid confusion, misunderstandings or potential misuse arising from differing interpretations between manufacturers and distributors, as well as the EU UDI issuing entities. This should...
EMA Questions & answers on Article 31 non-pharmacovigilance referral procedures
This guidance document addresses a number of questions which stakeholders, in particular the marketing authorisation holders (MAHs)/applicants, may have on an Article 31 non-pharmacovigilance referral procedure. It provides an overview of the European Medicines...
EXTENSION OF THE MDR TRANSITIONAL PERIOD AND REMOVAL OF THE ‘SELL OFF’ PERIODS
Q&A on practical aspects related to the implementation of Regulation (EU) 2023/607 amending Regulations (EU) 2017/745 and (EU) 2017/746 as regards the transitional provisions for certain medical devices and in vitro diagnostic medical devices REV. 2. JULY 2024
MHRA Updated guidance for the International Recognition Procedure (IRP)
📢Updated guidance for the International Recognition Procedure (IRP) is now available. As part of the MHRA commitment to continuously improving our regulatory processes, we've updated the IRP guidance to reflect several changes that will come into effect...
MDCG 2021-5 Rev. 1 .Guidance on standardisation for medical devices
This document aims to provide guidance on different aspects related to standards in the medical devices sector in support of the requirements laid down in the applicable EU legislation, taking into account its specificities
Qualification of novel methodologies (QoNM) for medicinal product development . Procedural Guidance to applicants
The EMA Qualification of Novel Methodologies (QoNM) provides a voluntary, scientific pathway to support development, evaluation and regulatory endorsement of innovative methodologies in specific contexts of use. In this guidance, the term ‘methodologies’ refers to...
FDA Guidance for Drug Development
The FDA has released a revised draft guidance (June 2026) on Master Protocols for Drug and Biological Product Development, reinforcing the growing role of innovative clinical trial designs in accelerating drug development.
Study supporting the monitoring of the availability of medical devices on the EU market. 2 July 2026
Survey results of the 20th NB survey (MDR/IVDR) with data status 28 February 2026 (small and medium dataset)
IMDRF Technical Framework for Artificial Intelligence Life Cycle Management” (IMDRF AIML WG N93 – April 2026)
Closing date: Friday, 10 July 2026 This document is intended for manufacturers that are developing AI-enabled medical devices to identify considerations as they make critical technical and governance choices. It is intended to provide considerations specific to...
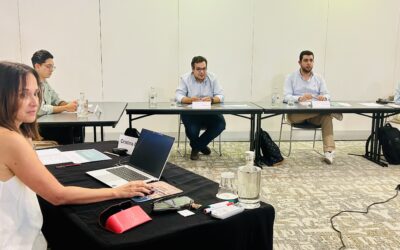
GMP CANÁBIS 2026 : “Formação conduzida com profundo domínio teórico e prático “
“Muito interessante em termos de conteúdos . Mas também na partilha de experiências . Muito enriquecedor “. NATECH No passado dia 30 de junho decorreu em Lisboa, a formação GMP CANÁBIS , bajo o lema : Visão integrada sobre os processos GACP (NOVA guideline...
MHRA Open consultation Draft rare disease therapies regulatory framework
This novel pathway also places significant emphasis on patient engagement and communication, ensuring that people affected by rare diseases are involved throughout development and that informed consent is an ongoing, not one‑off, process. The guidance also explains...
EU Delegated Acts on Well-Established Technologies (WET) under the MDR
The European Commission has published two delegated acts on Well-Established Technologies (WET) under the Medical Devices Regulation (EU) 2017/745 (MDR) in the Official Journal of the European Union. The newly published delegated regulations expand the list of WET...
CLINICAL TRIALS: EMA CTIS Sponsor Handbook
26 June 2026 EMA/186412/2021 Clinical Trials Transformation Sponsor handbook :Clinical Trial Information System (CTIS) user guidance on the sponsor’s workspace Version 6.3 The most Frequently Asked Questions (FAQs) on CTIS can be found in the Sponsor FAQs,...
Update to EU GMP Guidelines – Revised Annex 19 on Reference and Retention Samples
The European Commission has published a revised version of Annex 19 of EudraLex Volume 4. The update is focused on Section 9 of the annex which addresses reference and retention samples for parallel traded products. The revised annex includes expectations around the...
FARMACOVIGILÂNCIA nível avançado : Formação completa con casos práticos
“Formação bastante abrangente e completa. “ LAB EDOL No passado dia 23 de junho decorreu em Lisboa a formação GVP advanced : FARMACOVIGILÂNCIA nível avançado, desenhada para consolidar e atualizar conhecimentos e permitir o reforço dos sistemas...
FDA DRAFT GUIDANCE :Demonstrating Substantial Evidence of Effectiveness for Human Drug and Biological Products
This document is intended to provide guidance to applicants planning to submit new drug applications (NDAs), biologics license applications (BLAs), or supplements to NDAs or BLAs that require the demonstration of substantial evidence of effectiveness. This guidance...
European Biotech Act: Positioning EU as Biotech Powerhouse
10 things to know about the proposed European Biotech Act: Positioning EU as Biotech Powerhouse

CIOMS Glossary of ICH Terms and Definitions. Version 10
The CIOMS has released Version 10 of its Glossary of ICH Terms and Definitions, bringing together terminology from current International Council for Harmonisation (ICH) guidelines and Q&A documents. The glossary serves as a centralised reference for professionals...

Guidance on standardisation for medical devices Appendix: Transition to the ‘EU REP’ symbol in EN ISO 15223-1
The European Commission has published a new appendix to MDCG 2021-5 Rev. 1 – Guidance on standardisation for medical devices: 📄 Appendix: Transition to the “EU REP” symbol in EN ISO 15223-1 📅 Published: June 2026 This follows the publication in the...

MDCG Position Paper: Management of SS(C)P in EUDAMED after mandatory use
The Medical Device Coordination Group (MDCG) has published MDCG 2026-4: Management of SS(C)P in EUDAMED after Mandatory Use, providing guidance on how Summary of Safety and Clinical Performance (SSCP) and Summary of Safety and Performance (SSP) documents will be...

Commission Implementing Decision amending Implementing Decision (EU) 2021/1195 as regards the harmonised standard for symbols to be used with information to be supplied by the manufacturer
📢 Regulatory Update: Commission Implementing Decision (EU) 2026/1313 On 15 June 2026, the European Commission adopted Commission Implementing Decision (EU) 2026/1313, amending Implementing Decision (EU) 2021/1195 regarding the harmonised standard for symbols...
Publicado o Decreto-Lei n.º 118/2026 : Novos termos do SiNATS – Sistema Nacional de Avaliação de Tecnologias de Saúde (ATS)
Publicado o Decreto-Lei n.º 118/2026, de 17 de junho, que estabelece os novos termos do SiNATS - Sistema Nacional de Avaliação de Tecnologias de Saúde (ATS) O Decreto-Lei n.º 97/2015, de 1 de junho, instituiu, no âmbito do INFARMED - Autoridade Nacional do...
MDR Certification Process (including Pre-application, Application and Post Application phases) – Consensus document
The purpose of this consensus document is to describe in detail the pre-application, application processes through which manufacturers may apply to Notified Bodies (NBs) for the certification of medical devices under the regulation (EU) 2017/745 (MDR). The document...
Product Management Service (PMS) – Implementation of International Organization for Standardization (ISO) standards for the Identification of Medicinal Products (IDMP) in Europe Annex B to Chapter 1: Registration requirements to Public PMS API – Beta release
This document provides a step-by-step description of the registration process required to obtain access to the Public Product Management Service (PMS) Application Programming Interface (API) environment. The Public PMS API supports access to data managed within the...
4th progress report on the use of real-world evidence (RWE) in medicines regulation
A fourth report based on studies EMA conducted using real-world data between February 2025 and February 2026 is available. The report summarises the progress in integrating real-world evidence in regulatory decision-making. It contains a list of research topics and a...

ICH HARMONISED GUIDELINE GUIDELINE FOR GOOD CLINICAL PRACTICE E6(R3) Annex 2 .Final version
INTERNATIONAL COUNCIL FOR HARMONISATION OF TECHNICAL REQUIREMENTS FOR PHARMACEUTICALS FOR HUMAN USE HARMONISED GUIDELINE GUIDELINE FOR GOOD CLINICAL PRACTICE E6(R3) Annex 2. Final version Adopted on 03 June 2026

Team-NB Position Paper: MDR Certification Process (including Pre-application, Application and Post Application phases) – Consensus document
The purpose of this consensus document is to describe in detail the pre-application, application processes through which manufacturers may apply to Notified Bodies (NBs) for the certification of medical devices under the regulation (EU) 2017/745 (MDR). The document...
2025 AI Observatory report
Artificial Intelligence (AI) is fast evolving and is rapidly transforming the development and supervision of medicines. The EU AI Act entered into force in 2024 and represents a significant step towards ensuring that AI is developed and used in a trustworthy,...